Orgo Unit 3
 Call Kai
 Learn
 Practice Test
 Spaced Repetition
 Match
 Flashcards
 Knowt Play
1/66

There's no tags or description
Looks like no tags are added yet.
67 Terms
Preparation of Alkenes from Secondary and Tertiary Alcohols
Occurs through an E1 pathway involving a carbocation species, which often results in carbocation rearrangements and loss of stereospecificity. Driven by heating with strong acidic catalysts such as H₂SO₄ or H₃PO₄, or using Lewis acids (Al₂O₃)`
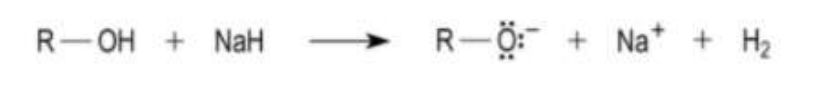
Formation of Alkoxides (R-O⁻)
Formed from the reaction of alcohols with stronger bases such as sodium hydride, NaH, which is a source of the hydride ion, H:
Formation of Mercaptides
Formed by the reaction of thiols (much more acidic than water or alcohols) with reaction with one equivalent of hydroxide or alkoxide OR
Thiols form insoluble mercaptides (sulfides) with many heavy-metal ions, such as Hg²⁺, Cu⁺, and Pb²⁺
Sulfonate Esters
Acidic hydrogen of a sulfonic acid is replaced by an alkyl or aryl group
**Have approximately the same reactivity as the corresponding alkyl bromides in substitution and elimination reactions

Preparation of Sulfonate Esters from Alcohols
Nucleophilic substitution reaction in which the oxygen of the alcohol displaces the chloride ion from tosyl chloride. The pyridine used as the solvent is a base.
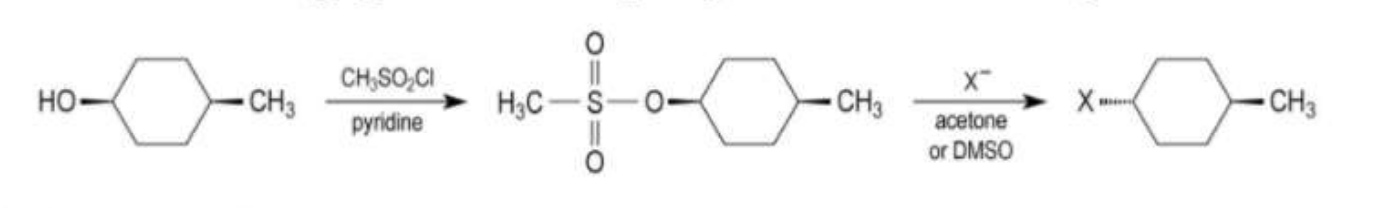
*The formation of sulfonate esters (tosylates/mesylates) perfectly retains the stereochemistry of the alcohol because the carbon-oxygen bond is never broken. However, the subsequent displacement of that ester via an SN2 pathway results in the inversion of stereochemistry

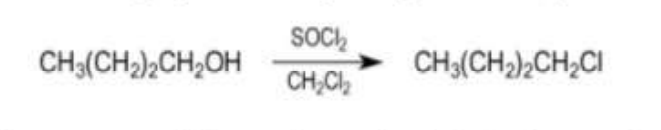
Preparation of Primary Alkyl Chlorides
Prepared from the reaction of primary alcohols with thionyl chloride (SOCl2) over CH2Cl2
Preparation of Primary Alkyl Bromides
Prepared from the reaction of primary alcohols with concentrated HBr or with Ph3PBr2

Preparation of Primary Alkyl Iodides
Prepared from the reaction of primary alcohols with concentrated HI (often provided by mixing KI and H3PO4)

Preparation of Primary Alkyl Halides using Sulfonate Esters
Works well for primary alkyl halides but requires two separate reactions: a) Formation of the sulfonate ester; b) Reaction of the ester with halide ion.

Mechanism of Primary Alkyl Halide Reactions
SN2…meaning alcohols with a significant amount of 𝛽-branching (alkyl substituents at the 𝛽-position) will react more slowly or not at all
Preparation of Secondary Alkyl Chlorides
Prepared from the reaction of secondary alcohols with thionyl chloride (SOCl2) → secondary alcohol has relatively little 𝛽-branching
*If the alcohol is hindered, use PPh3Cl2

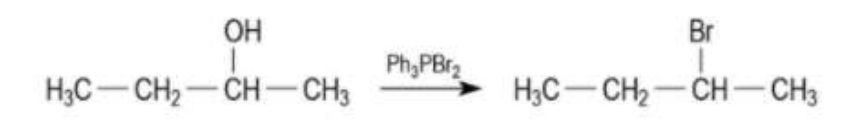
Preparation of Secondary Alkyl Bromides
Prepared from the reaction of secondary alcohols with Ph3PBr2
Preparation of Secondary Alkyl Halides using Sulfonate Esters
Prepared from the reaction of secondary alcohols with the sulfonate ester, which should be treated with the appropriate halide ion in a polar aprotic solvent (DMSO or acetone) to avoid rearrangements.
Mechanism of Secondary Alkyl Halide Reactions:
SN1: Treating an alcohol with strong acids like HCl, HBr, HI (generates a carbocation intermediate after the -OH group is protonated and leaves)
*Results in loss of stereochemistry
SN2: Treating an alcohol with SOCl3 or PPhBr2 (-OH is protonated as the nucleophile attacks the other side of the molecule)
*Results in inversion of stereochemistry
Preparation of Tertiary Alkyl Chlorides
Prepared from the reaction of tertiary alcohols with HCl under mild conditions
Preparation of Tertiary Alkyl Bromides
Prepared from the reaction of tertiary alcohols with HBr under mild conditions
Preparation of Tertiary Alkyl Halides using Sulfonate Esters
Do not occur (must occur through SN2)
Mechanism of Tertiary Alkyl Halide Reactions
SN1 (-OH is protonated as the nucleophile attacks the other side of the molecule)
Preparation of Alkenes from Alcohols
Prepared by the acid-catalyzed dehydration of alcohols
*The relative rates of alcohol dehydration are in the order: tertiary > secondary >> primary
E2 Reactions of Sulfonate Esters
Undergo these reactions with alkoxide bases (conjugate base of alcohol)
*This reaction is especially useful when the acidic conditions of alcohol dehydration lead to rearrangements or for primary alcohols in which dehydration is not an option

Oxidation of Primary and Secondary Alcohols to Aldehydes and Ketones:
Oxidized by reagents containing Cr(VI) (chromium in the +6 oxidation state) to give carbonyl compounds (C=O)
*The ɑ-carbon atom of an alcohol must bear a hydrogen atom for oxidation to an aldehyde or ketone to occur
*Primary alcohols react with Cr(VI) reagents to give aldehydes, but if water is present, aldehydes are further oxidized to carboxylic acids.

Forms of Chromium used in Oxidations
Chromate ion (H2CrO4), dichromate ion (KCr2O7), and chromic anhydride or chromium trioxide (CrO3)
Pyridine Chlorochromate (PCC)
Often used as an anhydrous Cr(VI) preparation for the oxidation of primary alcohols to aldehydes.
Mechanism of Oxidation of Primary and Secondary Alcohols to Aldehydes and Ketones
E2-like
Oxidation of Primary Alcohols to Carboxylic Acids
-Prepared from the reaction of the primary alcohols with aqueous solutions of Cr(VI), such as aqueous potassium dichromate (K2Cr2O7) in acid OR
-Primary alcohols can be oxidized to carboxylic acids with potassium permanganate (KMnO4) in basic solution, followed by acidification
*Because KMnO4 reacts with alkene double bonds, Cr(VI) is preferred for the oxidation of alcohols that contain double or triple bonds
*KMnO4 is not used for the oxidation of secondary alcohols to ketones because many ketones react further with the alkaline permanganate reagent.
Products of Oxidation of Thiols
The most commonly occurring oxidation products of thiols are disulfides and sulfonic acids
*Take place at sulfur, not carbon!
*When thiols and disulfides are present together in the same solution, an equilibrium among them is rapidly established.

Formation of Sulfonic Acids
Formed by vigorous oxidation of thiols or disulfides with KMnO4 or HNO3.

Formation of Disulfides
Formed by reaction of thiols with mild oxidants such as I2 in base or Br2 in CCl4
*Many thiols spontaneously oxidize to disulfides merely on standing in air (O2)!
*These reactions can be viewed as a series of SN2 reactions in which halogen and sulfur are the electrophiles that react with nucleophilic thiolate anions

Nucleophilic Substitution (Displacement) Reaction
A substitution because one group displaces another group, which departs.
The group that is displaced is called the leaving group (which is the nucleophile in the reverse reaction)
The nucleophile acts as a substituting group
What Makes a Good Leaving Group?
Gives the weakest bases as products (Strong conjugate acids)
Large, can stabilize the negative charge over multiple atoms/a large electron cloud
Polarizable
What Makes a Good Nucleophile?
Anionic
Low electronegativity
Large, polarizable
SN2 Reaction
Nucleophilic substitution reaction in which the leaving group is displaced from the electrophile in one concerted step
Typically carried out in polar protic solvents or polar aprotic solvents ((polar aprotic solvents in many cases accelerate SN2 reactions because they do not form hydrogen bonds to the nucleophile!!))
*Causes inversion of stereochemistry!
*Relative reactivity of carbons: Primary > Secondary >> Tertiary (due to steric hindrance)

SN1 Reaction
Nucleophilic substitution reaction in which the leaving group is displaced from the electrophile in two steps, forming a carbocation intermediate … the nucleophile is the solvent
Typically carried out in polar protic solvents
*Relative reactivity of carbons: Tertiary > Secondary >> Primary (due to unstable carbocation intermediate)

α-Hydrogen
Hydrogen atom attached to a carbon directly bound to the principal functional group

𝛽-Hydrogen
Hydrogen atom bound to the carbon directly adjacent to the ɑ-carbon (-carbon), positioned two carbons away from the principal functional group

Elimination Reaction
Reaction in which two or more groups are lost from within the same molecule
An elimination that involves loss of two groups from adjacent carbons is called a 𝛽-elimination (conceptually the reverse of an addition to an alkene)
E2 Reaction
𝛽-elimination reaction in which two bonds are broken, and a double bond is formed in one concerted step
Typically carried out in polar protic or polar aprotic solvents
*Typically occurs with anti-stereochemistry (hydrogen and leaving group are 180o apart)
*Relative reactivity of carbons: Tertiary > Secondary >> Primary

E1 Reaction
𝛽-elimination reaction in which the solvent acts as a weak base to remove the 𝛽-hydrogen, forming a carbocation intermediate
Typically carried out in polar protic solvents
*Relative reactivity of carbons: Tertiary > Secondary >> Primary (due to unstable carbocation intermediate)

Grignard Reagent
Organometallic compound of the form R-MgX, where X = Br. Cl, or I
Organolithium Reagent
Organometallic compounds of the form R-Li
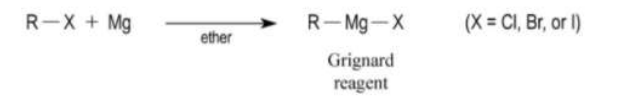
Formation of Grignard Reagents
Prepared from the reaction of alkyl or aryl halides with vigorously agitated Mg suspensions (under anhydrous conditions)
*Ethers are required as solvents!
Formation of Organolithium Reagents
Prepared from the reaction of alkyl halides with vigorously agitated Li suspensions (under anhydrous conditions)
*Hydrocarbons are required as solvents!

Reactions with Grignard and Organolithium Reagents
Organometallic compounds behave as strong Brønsted bases and react violently with acids, including water and alcohols, to give the conjugate-base hydroxide or alkoxides and the conjugate-acid hydrocarbon of the carbanion (protonalysis)
*Because carbon is more electronegative than either magnesium or lithium, the negative end of the carbon-metal bond is the carbon atom, essentially a carbanion. Carbanions are powerful Brønsted bases because their conjugate acids, the corresponding alkanes, are extremely weak acids.
Free-Radical Halogenation of Alkanes
Alkanes react with bromine and chlorine in the presence of heat or light in free-radical substitution reactions to give alkyl halides (follows the typical pattern of other free-radical chain reactions; it has initiation, propagation, and termination steps)
Initiated when a halogen molecule absorbs energy from heat or light and dissociates homolytically into halogen atoms.
*Halogenation with fluorine is violent, and halogenation with iodine does not occur.

Formation of Dihalocarbenes
Reaction of a haloform (like CHCl3) with base in the presence of an alkene yields a 1,1-dihalocyclopropane
*Concerted syn-addition reaction

Simmons-Smith Reaction
Method of preparing cyclopropane without halogen atoms by allowing alkenes to react with methylene iodide (CH2I2) in the presence of a zinc-copper couple
*Analogous to formation of a Grignard reagent
*Syn-addition

Williamson Ether Synthesis
Method of preparing an ether in which the conjugate base of an alcohol (or thiol) reacts as a nucleophile with an unhindered alkyl halide or sulfonate ester to form an ether (or sulfide)
*SN2 mechanism! (Tertiary and many secondary alkyl halides cannot be used because the E2 (elimination) reaction occurs instead → the preferred route always uses the less sterically hindered alkyl halide)

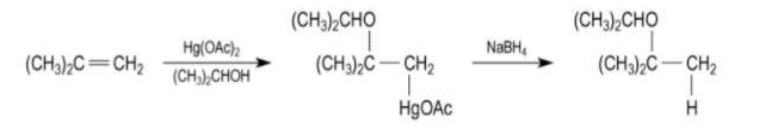
Alkoxymercuration-Reduction of Alkenes
Essentially oxymercuration-reduction carried out in an alcohol solvent instead of water (an alcohol instead of water is the nucleophile that attacks the mercurinium ion intermediate)
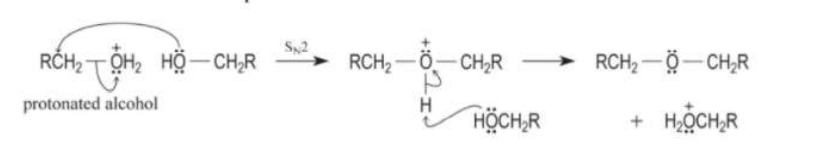
Ethers from Primary Alcohol Dehydration
Method of preparing symmetrical ethers by dehydrating primary alcohols with strong acid and heat
*Secondary and tertiary alcohols cannot be used because they undergo dehydration to alkenes.
*SN2 reaction in which one alcohol displaces water from another molecule of protonated alcohol.
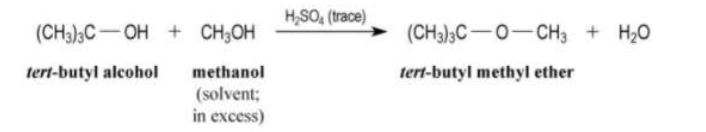
Ethers from Tertiary Alcohol Dehydration
Method of preparing unsymmetrical ethers by treating them with dilute strong acid in alcohol solvents
*SN1 solvolysis reaction in which the protonated alcohol loses water to give a relatively stable carbocation that is attacked by the solvent to give the product.
Ethers from Alkene Dehydration
Treatment of an alkene with a large excess of alcohol in the presence of an acid catalyst gives an ether…carbocation intermediate.

Oxidation of Alkenes with Peroxycarboxylic Acids
Method of preparing epoxides by directly oxidizing alkenes with peroxycarboxylic acids (R-COOH)
*Concerted addition reaction, similar to that of Br2 addition to alkenes
*Stereospecific…retention of the alkene stereochemistry

Epoxides from the Treatment of Halohydrins with Base
Intramolecular variation of the Williamson ether synthesis in which the alcohol and the alkyl halide are part of the same molecule
Alkoxide anion (NaOH + alcohol) displaces the halide ion from the neighboring carbon
*SN2 reaction
*Halohydrins derived from cyclic compounds must be able to assume the required anti relationship through a conformational change!

Alkene Reaction with Diethyl Tartrate (DET):
(+)-DET adds syn- from the bottom to form an epoxide
(-)-DET adds syn- from the top to form an epoxide

Preparation of Glycols by the Acid-Catalyzed Reaction of Water with Epoxides
Converts an epoxide into a 1,2-diol (glycol) with anti-stereochemistry (trans-diol configuration) at the more substituted carbon atom

Preparation of Glycols by the Oxidation of Alkenes with OsO4
A method of preparing glycols by treatment with OsO4 (OsO4 is in a high oxidation state. Results in a concerted cycloaddition and gives an intermediate called an osmate ester.) A glycol is formed when the cyclic osmate ester is treated with water (and a mild reducing agent, NaHSO3)
*By including an amine oxide in the reaction mixture, only catalytic amounts of OsO4 are required!
*Stereospecific syn-addition

Preparation of Glycols by the Oxidation of Alkenes with KMnO4
Stereospecific syn-addition, usually under aqueous alkaline conditions.
*When oxidation occurs, the brilliant purple color of the ion is replaced by a murky brown precipitate of magnesium dioxide (Baeyer test)
*Yields are low in many cases because over-oxidation occurs …

Ether Cleavage
The C-O bonds of ethers react under acidic conditions (HI or HBr) to give alcohols and alkyl halides
*Ethers containing only methyl and/or primary alkyl groups require strong acid and relatively harsh conditions
*Secondary and tertiary ethers cleave more readily than primary or methyl ethers because the SN1 mechanism intervenes and carbocation intermediates can be formed

Cleavage of Ethers with Primary/Methyl Alkyl Groups
Involves protonation of the ether oxygen, followed by a substitution reaction of the protonated ether by the iodide or bromide ion (a good nucleophile)
*SN2 reaction to form an alkyl halide, which liberates an alcohol as a leaving group

Cleavage of Ethers with Tertiary Alkyl Groups
Involves protonation of the ether oxygen, followed by formation of a carbocation by loss of the alcohol leaving group, and finally a Lewis acid—base association reaction of the carbocation with the halide ion.
*Can be done under milder conditions
*Because the SN1 reaction is faster than competing SN2 processes, none of the primary alkyl halides are formed.

Epoxide Ring Opening Under Basic Conditions
Involves backside substitution by the nucleophile (alcohol anion) at the less substituted epoxide carbon
*Essentially an SN2 reaction where the epoxide oxygen is the leaving group → Inversion of configuration is observed if attack occurs at a stereocenter!

Epoxide Ring Opening Under Acidic Conditions
Involves substitution at the more substituted epoxide carbon
*The more substituted carbon has more delta+ character
*Inversion of configuration is observed if attack occurs at a stereocenter

Preparation of Glycols by Acid-Catalyzed Epoxide Hydrolysis
This reaction occurs by backside substitution of water on the protonated epoxide
When combined with epoxide formation from an alkene, gives glycols with different stereochemistry from that obtained from the OsO4 reaction with alkenes

Reaction of Ethylene Oxide and Grignard Reagents
Carbanionic C-Mg carbon acts nucleophilically toward the epoxide ring. Magnesium functions as a Lewis acid, binding oxygen to enhance leaving-group ability. After the Grignard reagent has reacted, the alkoxide is converted into the alcohol product in a separate step by the addition of water or dilute acid.
*Provides a method for the formation of carbon-carbon bonds!

Oxidative Cleavage of Glycols
Method of cleaving the C-C bond between the -OH groups of a vicinal glycol by periodic acid
*Takes place through a cyclic periodate ester intermediate, which spontaneously breaks down by a cyclic flow of electrons in which the iodine accepts an electron pair.
*A glycol that cannot form a cyclic ester intermediate is not cleaved by periodic acid!

Oxidation of Ethers and Sulfides
Ethers are relatively inert towards oxidizing conditions
Sulfides are readily oxidized to sulfoxides and sulfones.
→ Sulfoxides and sulfones can be prepared by the direct oxidation of sulfides with one and two equivalents, respectively, of hydrogen peroxide HOOH
*Other common oxidizing agents, such as KMnO4, and peroxyacids readily oxidize sulfides

Reactions of Oxonium and Sulfonium Salts
If the acidic hydrogen of a protonated ether is replaced with an alkyl group, the resulting compound is called an oxonium salt; the sulfur analog of an oxonium salt is a sulfonium salt. Oxonium and sulfonium salts react with nucleophiles in SN2 reactions.
*Oxonium salts are usually isolated as BF4 salts
*Sulfonium salts are considerably less reactive than the corresponding alkyl chlorides in SN2 reactions
